FDA 510k Consultants for Medical Devices and IVDs
MedDeviceCorp's FDA 510k consultants take your device from classification to 510k clearance. We find the right predicate, plan your testing, prepare the eSTAR, file it with the FDA, answer every FDA question on your behalf and act as your US Agent. You keep control of the decisions; we carry the regulatory workload.
Our 510k consulting team has over 10 years of technical and scientific experience with US FDA submissions for non-invasive, invasive and implantable devices and in vitro diagnostics, supporting manufacturers from offices in India, the USA and the UK.
Reviewed by Beena P, Manager – USFDA Regulatory & Operations, 15+ years in regulatory consulting · Last updated: October 2026
Get a 510k QuoteSee 510k Consulting FeesWhy Work With 510k Consultants?
A 510(k) looks simple on paper: show that your device is substantially equivalent to a legally marketed predicate. In practice, most delays come from a weak predicate, testing that does not match FDA guidance, or slow answers to the FDA's Additional Information (AI) request. Experienced 510k consultants prevent those problems before the file reaches the FDA.
Right first time
A complete, consistent eSTAR passes the FDA technical screening and reduces the questions you get during substantive review.
Lower testing cost
We define only the tests your predicate, device type and FDA guidance actually call for, so you avoid paying for studies the FDA will not need.
Faster responses
We handle FDA correspondence and AI responses within the 180-day window, so your clearance is not lost to a missed deadline.
One point of contact
Consultant, submitter correspondent and US Agent in one team, from first assessment until you receive your K number.
Our 510k Consulting Services
Choose the complete 510(k) package or only the services you need. Our FDA 510k consultants and 510k submission consultants cover every stage:
Gap assessment
Product analysis and review of your technical documentation against FDA requirements, guidance and recognized standards.
Classification and predicate
Product code, regulation number and class, 510(k) type (Traditional, Special or Abbreviated) and predicate device search.
Test strategy
Performance, biocompatibility, electrical safety, software and sterilization test requirements, with review of laboratory protocols and reports.
Substantial equivalence
Technical comparison of your device with the predicate, with a clear justification for every difference.
Labeling review
Review and modification of labels, IFU and device description to match your indications for use and 21 CFR 801 or 809.
FDA Pre-Submission
Q-Sub preparation and FDA meeting support to agree your testing plan before you commit budget.
eSTAR preparation
We draft, edit, review and compile the complete 510(k) in the FDA eSTAR template, ready for submission through the CDRH Portal.
Submission and FDA follow-up
User fee coordination, submission, and acting as your correspondent with the FDA until the decision.
AI response
Responses to FDA Additional Information requests in the shortest possible time frame, with you supplying any new data.
US Agent
For foreign manufacturers we act as US Agent and pass on every FDA communication immediately. US FDA Agent
Registration and listing
Establishment registration and device listing with your 510(k) number after clearance. US FDA Registration
Device changes
Assessment of design, labeling or software changes and Special 510(k) submissions when a new clearance is needed.
Devices Our 510k Consultants Support
Non-invasive devices
Monitoring equipment, diagnostic instruments, physiotherapy and rehabilitation devices, wound dressings and similar products.
Invasive devices
Catheters, surgical instruments, needles and other devices that enter the body through an orifice or the skin.
Implants
Orthopedic, dental and other implantable devices that need detailed biocompatibility, mechanical and sterility evidence.
IVDs
In vitro diagnostic devices needing analytical and clinical performance data, including point-of-care tests.
Devices with software
Electromedical devices and device software functions needing software documentation and, for cyber devices, cybersecurity content.
How Our 510k Consulting Process Works
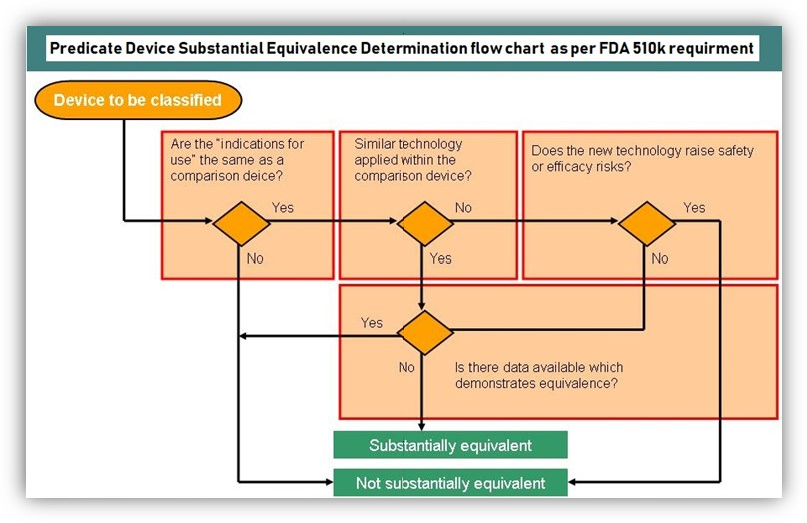
Our 510k consultants start with the same decision the FDA reviewer makes: is your device substantially equivalent to the predicate? The flowchart shows how that decision is reached.
- Confirm the device and its regulatory status, then identify the product code, regulation number and class.
- Agree the predicate with you: the 510(k)-cleared device with the same intended use and closest technology.
- Select the 510(k) type: Traditional, Abbreviated or Special.
- Gap assessment and test plan, including preclinical studies and testing at external accredited laboratories.
- FDA Pre-Submission where needed, to confirm the testing strategy.
- Prepare the 510(k) in eSTAR and update it with Pre-Sub feedback and final test reports.
- US Agent appointment and FDA user fee payment.
- Submit the eSTAR to the FDA through the CDRH Portal.
- Follow up with the FDA on your behalf during technical screening and substantive review.
- Respond to FDA review comments, modify the 510(k) and resubmit.
- Receive the 510(k) number and clearance letter.
- Register the establishment and list the device with the 510(k) number.
- Start selling in the USA, with a QMSR-compliant quality system in place.
For the FDA's side of the process, review clock and timelines, see our FDA 510k submission guide.
Who Does What: You and Your 510k Consultants
A 510(k) is a joint effort. This is how responsibilities are shared in a complete package.
| Activity | Client | MedDeviceCorp |
|---|---|---|
| Select device and models for the US market | Yes | – |
| Predicate identification and 510(k) vs De Novo or PMA decision | Yes | Yes |
| Product code, class and applicable guidance | – | Yes |
| Biocompatibility and test requirement identification | – | Yes |
| Sending samples to laboratories | Yes | – |
| Review of risk analysis, test protocols, labels, IFU and shelf life | Yes | Yes |
| Pre-Submission | Yes | Yes |
| Drafting and compiling the eSTAR | – | Yes |
| FDA user fee payment | Yes | – |
| Submission and FDA correspondence | – | Yes |
| Additional data for FDA questions | Yes | Yes |
| Modification and resubmission of the 510(k) | – | Yes |
What You Receive
- Regulatory strategy with product code, regulation number, class and 510(k) type
- Predicate analysis and substantial equivalence comparison
- Test plan listing the standards and FDA guidance for your device
- Pre-Submission package and FDA feedback (where included)
- Complete 510(k) in eSTAR format, including the 510(k) summary
- Submission confirmation and FDA correspondence log
- Responses to FDA Additional Information requests
- 510(k) clearance letter support, followed by registration and listing
All documentation is managed on a secure cloud server, and communication is in English by email, phone, video call or WhatsApp.
510k Submission Consultants: Problems We Fix
Many manufacturers come to our 510k submission consultants after a first attempt has stalled. These are the issues we see most often, and how we resolve them:
| Problem | What happens | How we fix it |
|---|---|---|
| Weak or mismatched predicate | Questions on intended use or technology, or a Not Substantially Equivalent decision | Re-run the predicate search by product code and indications, and rebuild the comparison |
| Incomplete eSTAR | The file is put on hold at technical screening | Complete every eSTAR section with consistent device names, models and data |
| Testing gaps | AI requests for missing biocompatibility endpoints, bench tests or shelf-life data | Map tests to FDA guidance and recognized standards before the file goes in |
| Software and cybersecurity | Requests for documentation at the right level, or Section 524B cybersecurity content | Prepare software and cybersecurity documentation to current FDA guidance |
| Labeling inconsistencies | Indications in the IFU do not match the indications for use statement | Align labels, IFU and the 510(k) so every claim is supported |
| Missed AI deadline | The 510(k) is considered withdrawn after 180 days and must be resubmitted with a new fee | Plan the response from day one and manage the timeline with you |
In-House Team or FDA 510k Consultants?
Some manufacturers build an internal regulatory team; others hire FDA 510k consultants for each submission. Many do both, with consultants supporting the internal team on complex files.
| Factor | In-house only | With 510k consultants |
|---|---|---|
| FDA submission experience | Depends on your hires; often limited to one device type | Experience across many device types, panels and FDA reviewers |
| Cost | Fixed salaries all year | Fixed fee per 510(k), paid in stages |
| Speed to start | Hiring and training time | Work starts as soon as you sign |
| US Agent and US-hours contact | Needs a separate provider | Included with MedDeviceCorp |
| Knowledge of current FDA practice | Must be tracked internally | eSTAR, guidance updates and review trends tracked daily |
If you already have a regulatory team, our partial 510k consulting services, such as a detailed 510(k) review or Pre-Submission support, add an expert second check without replacing your people.
510k Consulting Fees
Our 510k consulting fees are fixed by device type, so you know the cost before you start. FDA user fees, laboratory testing and taxes are separate. For current FDA fees, see FDA 510k submission fees.
Option 1: Complete 510k package (payable to MedDeviceCorp)
Covers 510k consulting, preparation, Pre-Submission, final submission and US Agent.
| Type of device | Guidance fee | 510(k) file preparation | Submission fee |
|---|---|---|---|
| Non-invasive | 8,000 – 9,000 USD | 6,000 – 7,000 USD | 500 USD |
| Invasive | 10,000 – 11,000 USD | 8,000 – 9,000 USD | 500 USD |
| Implant | 12,000 – 13,000 USD | 10,000 – 11,000 USD | 500 USD |
Option 2: Partial 510k services (payable to MedDeviceCorp)
| Service | Cost | Remarks |
|---|---|---|
| US FDA 510(k) detailed review | 6,500 – 8,500 USD per 510(k) | Device code, regulation number and accurate predicate details must be provided by the client. Timeline 30 working days. |
| US FDA 510(k) quick review + FDA Pre-Submission* | 6,000 – 7,500 USD per 510(k) | Expected timeline 90 to 120 days. US Agent fees included. No FDA fee for the Pre-Submission. |
| US FDA 510(k) Pre-Submission | 3,500 USD per 510(k) | US Agent fees included. Expected timeline 90 days. No FDA fee. |
| US FDA 510(k) Pre-Submission + 510k submission | 6,500 USD per 510(k) | Pre-Submission about 90 days, 510(k) review about 120 days. FDA 510(k) review fee applies. US Agent fees included. |
* 510(k) submission can be added to this service for an additional 1,500 USD.
Important information
- Country-specific government taxes are additional.
- Onsite consultant visits are available on request; travel and boarding are invoiced at actual cost.
- Project cost is paid in 4 instalments based on project progress.
- FDA user fees are paid by the client directly to the FDA.
How Long Does 510k Consulting Take?
| Stage | Typical duration |
|---|---|
| 510(k) preparation | 60 to 90 days |
| FDA Pre-Submission (optional) | About 90 days |
| FDA review cycles to clearance | 140 to 180 calendar days |
Laboratory lead times for biocompatibility, electrical safety and other testing can extend preparation. See the full phase-by-phase plan in our 510k submission timeline.
How to Choose a 510k Consultant
Whether you hire us or another firm, check these points before you sign:
- Device experience: ask for 510(k) experience with your device type, product code or FDA review panel.
- Ownership of the file: the 510k consultant should write the submission, not just review what you draft.
- Clear scope and fees: fixed fees by stage, with FDA fees, testing and taxes shown separately.
- Current knowledge: eSTAR, cybersecurity under Section 524B, software documentation and the QMSR.
- FDA correspondence: the consultant should act as correspondent and handle AI responses, not hand them back to you.
- US Agent and registration: having these in the same team avoids gaps after clearance.
- Time zone coverage: a team that can work in your hours and in US hours.
FDA Review or Third Party Review?
Most 510(k)s are reviewed directly by the FDA. For eligible device types, your 510(k) can instead go to an FDA-accredited organization under the 510(k) Third Party Review Program, which reviews the file and sends its recommendation to the FDA. There is no FDA user fee for a third party review, but the organization charges its own fee. Our 510k consultants check whether your product code is eligible and which route is faster and more cost-effective for you.
510k Clearance and Your Quality System
The FDA does not inspect your facility as part of 510(k) review. But once you register and sell in the US, the FDA can inspect at any time. Since 2 February 2026 the Quality Management System Regulation (21 CFR 820) incorporates ISO 13485:2016, so manufacturers not exempt from 21 CFR 820 must have a compliant system in place. A failed inspection can lead to a warning letter and products being withdrawn from the market.
We can prepare your quality system alongside your 510(k): FDA QSR Certification and ISO 13485 consulting.
US FDA 510k Consulting From India, the USA and the UK
MedDeviceCorp works with manufacturers worldwide from offices in India, the USA and the UK. Our team works across time zones, so we can meet you in your working hours and still respond to FDA reviewers during US hours. Many clients combine their US FDA 510k consulting with EU and UK market access, so one technical file supports several submissions.
Related Services
EU market
Testing
Learn more
FDA 510k Consultants FAQs
What do FDA 510k consultants do?
They plan and prepare your 510(k): classification, predicate selection, test strategy, eSTAR preparation, submission, FDA correspondence and responses to FDA questions until clearance.
Do I need a 510k consultant?
It is not mandatory, but most manufacturers without in-house FDA experience use one to avoid technical screening holds, extra testing and delays.
How much do 510k consultants charge?
MedDeviceCorp's complete package ranges from about 14,500 USD for a non-invasive device to about 24,500 USD for an implant, plus FDA fees, testing and taxes. Partial services start at 3,500 USD.
Are FDA user fees included in your 510k consulting fees?
No. FDA user fees are paid directly by you to the FDA. Our fees cover consulting, preparation, submission and US Agent services.
How long does a 510(k) take with a consultant?
Preparation usually takes 60 to 90 days, a Pre-Submission about 90 days, and FDA review cycles to clearance 140 to 180 calendar days.
Will you write the 510(k) or only review it?
In the complete package we draft, edit, review and compile the full 510(k). Partial services include detailed or quick reviews of a file you have prepared.
Can you help find a predicate device?
Yes. We search the FDA databases for cleared devices with the same product code and intended use and agree the best predicate with you.
Do you arrange testing?
We define the tests needed and review laboratory protocols and reports. You send samples to accredited laboratories, and testing costs are paid to the laboratory.
Do you act as US Agent?
Yes. US Agent services are included in our complete package and in our Pre-Submission services for foreign manufacturers.
Can you respond to FDA Additional Information requests?
Yes. We act as your correspondent, prepare the response and resubmit, with you supplying any new test data needed.
Do you handle IVD 510(k)s?
Yes. We support IVD submissions, including analytical and clinical performance data requirements.
Can you help with a Special 510(k) for a device change?
Yes. We assess whether your change needs a new 510(k) and prepare a Special or Traditional 510(k) as appropriate.
Is a Pre-Submission included?
The complete package includes a Pre-Submission. It is also available as a separate service at 3,500 USD per 510(k).
How are payments made?
Project fees are split into 4 instalments linked to project progress.
Do you visit our facility?
Onsite visits are available on request, with travel and boarding invoiced at actual cost.
Does the FDA inspect before 510k clearance?
No, but the FDA can inspect after you start selling, so you need a QMSR-compliant quality system.
Can you register our establishment after clearance?
Yes. We handle establishment registration and device listing with your 510(k) number.
Is 510k clearance the same as FDA approval?
No. 510(k) devices are cleared, not approved. FDA approval applies to PMA devices.
Which countries do you support?
We support manufacturers worldwide from our offices in India, the USA and the UK.
How do I get a quote?
Send your device details through our contact page and our 510k consultants will reply with a detailed estimate.
Talk to Our FDA 510k Consultants
Share your device details and we will confirm the pathway, predicate, timeline and fixed fee for your 510(k).
Contact UsRead the 510k Submission Guide
Our Social Activities